China’s top pharma suppliers work in a field where a batch of tablets must be more than consistent in color and shape. Its ingredients, manufacturing records, stability data, and clinical evidence all matter. Why is drug approval process strict? Because a medicine’s benefits must be supported by reliable evidence, while risks need careful assessment before patients depend on it. Robert Califf, a physician and former FDA commissioner, has emphasized the need for evidence that medicines are safe and effective. That is the core standard. It sounds simple, but applying it can take years.
Reviewers examine details that may seem small: how an active ingredient is produced, whether a tablet dissolves as expected, and how side effects compare across study groups. Suppliers therefore need documented quality controls, traceable materials, and dependable data—not just a polished product sample. For manufacturers in China seeking international partnerships, clear records and consistent production can make review more credible. They do not guarantee approval. Nor should they. Strict review can delay access and raise development costs, and no process can remove every uncertainty. That deserves honest reflection. A decision is only as strong as the evidence behind it, and the evidence is never perfect. The goal is not paperwork for its own sake. It is a careful judgment that helps protect patients while allowing well-supported treatments to reach them.
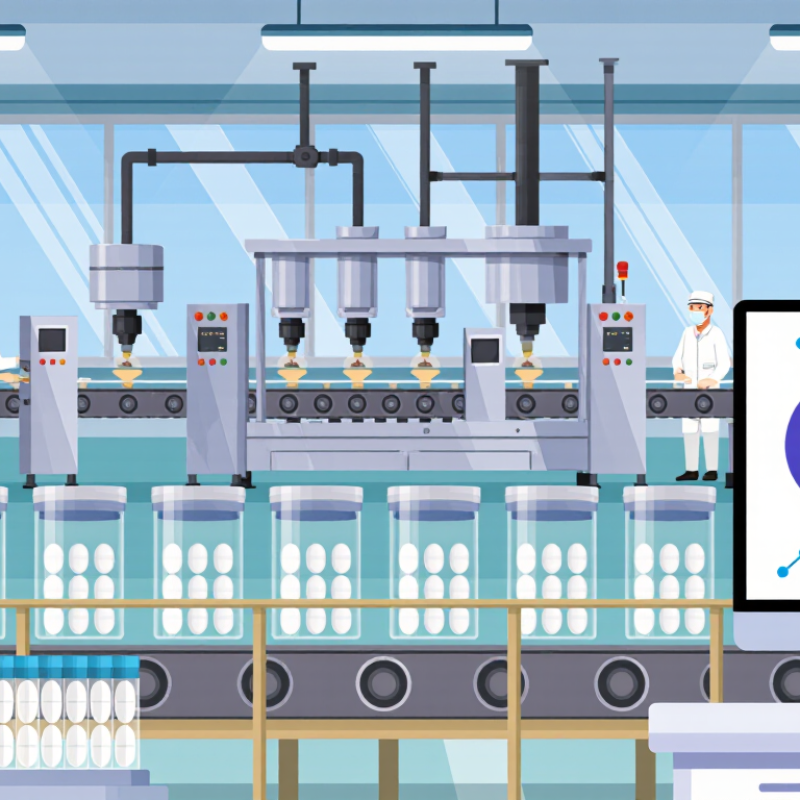
China’s drug approval system is strict because safety evidence must survive several regulatory checks. The framework is built around the Drug Administration Law, Good Clinical Practice, Good Manufacturing Practice, and the national pharmacopoeia. These rules cover research, clinical trials, production, distribution, and post-market monitoring.
The National Medical Products Administration is the main approval authority. Its Center for Drug Evaluation reviews clinical data, quality studies, and benefit-risk assessments.
Provincial medical products administrations inspect manufacturing sites and supervise local compliance. Ethics committees examine trial protocols and participant protection. Customs and health authorities may also support specific import, inspection, or public health procedures.
A typical application includes stability results, batch records, manufacturing controls, and clinical evidence. Reviewers may question a small detail, such as an unexplained laboratory variation or incomplete data traceability. That level of scrutiny can extend timelines, but it helps reveal risks before wider use. In practice, procedures are not always perfectly predictable. Requirements may be clarified during review, and applicants sometimes need to revise documents repeatedly.
This is frustrating.
For overseas suppliers, accurate Chinese-language documentation, consistent quality systems, and early regulatory planning are essential. A supplier should prepare for on-site inspections, sampling checks, and continuing pharmacovigilance after approval. Approval is not the final checkpoint. Safety reporting continues throughout the product’s lifecycle.
China Top Pharma Suppliers: Why Is Drug Approval So Strict?
New drug approval begins long before human testing. Researchers must study pharmacology, absorption, metabolism, and possible toxicity. Laboratory work often follows Good Laboratory Practice standards. Test results should be traceable, repeatable, and supported by clear records. Animal studies can reveal organ damage, immune reactions, or unsafe dose ranges. They cannot predict every human response. That limitation matters.
Clinical trials then examine whether a candidate is safe and useful for patients. Phase I usually evaluates tolerability, side effects, and drug behavior in the body. Phase II explores dosing and early evidence of benefit. Phase III involves larger patient groups and compares outcomes through defined endpoints. Investigators must follow approved protocols, obtain informed consent, and report serious adverse events quickly. Independent review helps protect participants and reduce biased decisions.
Manufacturers and suppliers also need reliable chemistry, manufacturing, and quality-control data. Each batch should meet identity, strength, purity, and stability requirements. Regulators examine trial records, production facilities, statistical methods, and risk-management plans. The process can feel slow, especially when promising treatments address urgent needs. Still, speed cannot replace evidence.
No study is flawless. Real-world patients may have conditions that trials did not fully represent. For this reason, ongoing safety monitoring remains essential after approval. Careful review protects patients, supports responsible suppliers, and builds trust in new medicines.
Drug approval is strict because a polished file cannot replace reliable evidence. For Chinese pharmaceutical suppliers, safety evidence begins with toxicology, dose selection, and adverse-event monitoring. Regulators examine trial protocols, patient records, and statistical methods. The FDA’s 2023 New Drug Therapy Approvals report recorded 55 novel drug approvals, showing how demanding the final review remains. Small data gaps can delay an otherwise promising product.
Effectiveness requires more than positive laboratory results. Controlled clinical trials must demonstrate meaningful benefits, not only numerical changes. The IQVIA Institute’s Global Trends in R&D 2024 reported more than 21,000 medicines in the global pipeline. Competition for approval is intense. Reviewers question weak control groups, short follow-up periods, and missing patient diversity. Sometimes, supplier teams overvalue a successful endpoint. That is understandable, but it can hide practical risks.
Product quality is equally important. Manufacturing sites need validated processes, traceable raw materials, stable storage conditions, and accurate batch records. ICH quality guidance expects suppliers to control impurities and process variation throughout production. A single unexplained deviation may trigger additional testing. No dossier is perfect. Human mistakes happen. The real test is whether the supplier detects, documents, and corrects them before patients face the consequence.
China Top Pharma Suppliers: Why Is Drug Approval So Strict?
Manufacturing Inspections and Good Manufacturing Practice Compliance
Drug approval is strict because quality cannot be judged by appearance alone. Inspectors examine whether a manufacturer can produce safe, consistent batches over time. They review batch records, raw material testing, equipment maintenance, and laboratory controls. A clean production room is not enough.
During a manufacturing inspection, small details can matter. Inspectors may check temperature logs, cleaning records, water-system tests, and employee training files. They also compare electronic data with paper records. Missing signatures or unexplained changes may raise serious questions. Traceability must work from incoming materials to final shipment.
GMP compliance requires more than written procedures. Workers must understand each step and follow it consistently. When a deviation occurs, the company should investigate its real cause. Corrective actions must be measurable, documented, and checked later. Weak explanations create weak confidence.
No system is flawless. Even experienced facilities can overlook a repeated equipment issue or a delayed record. That is why internal audits, risk reviews, and independent oversight remain important. In practice, reliable suppliers welcome difficult questions. They can show evidence, explain limitations, and improve without hiding uncomfortable findings. Strict approval protects patients, but it also tests whether manufacturing discipline is genuine.
Drug approval is strict because manufacturers must control the entire production system, not only test the finished product. The chart shows the number of active regulatory sections in each subpart of U.S. FDA 21 CFR Part 211, the Current Good Manufacturing Practice requirements for finished pharmaceuticals. These areas are commonly reviewed during manufacturing inspections, including personnel, facilities, equipment, production controls, laboratory testing, records, and distribution.
Source: U.S. Electronic Code of Federal Regulations, 21 CFR Part 211, Current Good Manufacturing Practice for Finished Pharmaceuticals. Section counts reflect the active numbered sections listed under each subpart.
How Strict Approval Rules Shape Chinese Pharmaceutical Suppliers
China’s strict drug approval rules reshape how pharmaceutical suppliers operate every day. The National Medical Products Administration reviews clinical evidence, manufacturing controls, and product quality before approval. A supplier cannot rely on a polished sample alone. Inspectors may examine batch records, laboratory methods, equipment maintenance, and staff training.
In practical supplier audits, small details often matter. A clean production room is not enough. Temperature logs must be complete, test results must be traceable, and deviations need documented investigation. Suppliers also need validated processes, qualified raw materials, and reliable pharmacovigilance systems after market entry. These requirements increase costs, but they reduce risks linked to contamination, unstable formulations, and inconsistent batches.
The rules also change supplier selection. Buyers increasingly assess regulatory history, inspection readiness, and data integrity before discussing price. A factory with modern equipment may still lose trust if records appear incomplete. This process can feel slow. Sometimes, it is frustrating. Yet strict review encourages suppliers to invest in trained quality teams and stronger internal controls. The system is not perfectly smooth, either; approval timelines can vary with product complexity and review questions. Suppliers that communicate clearly, correct weaknesses, and maintain evidence over time usually build more dependable partnerships.
| Regulatory area | Verified rule or framework | How it shapes pharmaceutical suppliers | Useful buyer checks |
|---|---|---|---|
| Drug registration | China’s Drug Registration Regulation, effective July 1, 2020, sets requirements for clinical-trial applications, marketing authorization, and post-approval changes. | Suppliers must support product-specific documentation and keep manufacturing and quality information consistent with the approved dossier. | Check the product’s registration status, approved specifications, and whether proposed changes have the required regulatory handling. |
| Clinical-trial review | For a clinical-trial application, the review period is 60 working days from acceptance; if the applicant receives no negative or questioning notice within that period, the application is considered approved under the applicable procedure. | Contract manufacturers and research-service providers may need to demonstrate controlled processes, reliable records, and readiness for inspections or audits. | Confirm the scope of any trial-related services and review relevant quality agreements, records, and inspection history. |
| Good Manufacturing Practice | China’s Drug Administration Law requires drug manufacturers to comply with Good Manufacturing Practice (GMP) and maintain a quality-management system covering manufacturing. | Production sites need controlled procedures for personnel, facilities, materials, manufacturing, testing, deviations, and records—not just finished-product testing. | Review site and product scope, audit findings, batch documentation, deviation controls, and corrective-action records. |
| Marketing Authorization Holder (MAH) | China’s nationwide MAH system makes the authorization holder responsible for the quality, safety, and efficacy of its drugs, including when manufacturing is outsourced. | Third-party manufacturers must operate within clearly defined responsibilities and provide the data and cooperation needed by the authorization holder. | Clarify responsibilities for release, change control, complaints, recalls, audits, and regulatory reporting in written agreements. |
| Generic-drug quality | China’s generic-drug consistency evaluation policy requires specified generic products to demonstrate consistency with the reference product in quality and, where applicable, therapeutic effect. | Suppliers may need robust formulation, analytical, process-validation, and bioequivalence support for products within the policy’s scope. | For relevant products, verify evaluation status and the supporting evidence applicable to that specific dosage form and product. |
| Pharmacovigilance | Marketing authorization holders in China must establish a pharmacovigilance system and collect, assess, and report adverse drug reaction information in accordance with applicable requirements. | Suppliers need procedures to promptly pass on relevant quality complaints and safety information to the responsible authorization holder. | Check complaint escalation timelines, safety-information responsibilities, traceability, and recall support. |
| Inspection and traceability | Regulators can conduct inspections and take risk-based supervisory measures. Drug manufacturers and authorization holders must maintain records and support the oversight of drug quality. | Suppliers are expected to maintain retrievable records, trace materials and batches, investigate problems, and address identified deficiencies. | Assess batch traceability, data integrity controls, inspection readiness, and documented corrective and preventive actions. |
| Note: Regulatory requirements depend on the product, activity, and applicable rules. Verify current requirements and product-specific status with official Chinese regulatory sources before making procurement or compliance decisions. | |||
Regulators need evidence of safety, quality, and possible patient benefit. Laboratory results alone are not enough. Human risks can remain uncertain.
Researchers study pharmacology, absorption, metabolism, and toxicity. Animal studies may reveal organ damage or unsafe dose ranges. They cannot predict every human reaction.
Phase I examines tolerability, side effects, and drug behavior. Phase II explores dosage and early benefits. Phase III compares outcomes in larger patient groups.
Investigators follow approved protocols and obtain informed consent. They report serious adverse events quickly. Independent review helps limit biased decisions.
Regulators review batch records, raw material tests, equipment maintenance, and laboratory controls. Each batch should meet identity, strength, purity, and stability requirements.
Inspectors may check temperature logs, cleaning records, water tests, and training files. They compare electronic data with paper records. Small gaps matter.
Workers must understand procedures and follow them consistently. Deviations need root-cause investigations and measurable corrective actions. Weak explanations reduce confidence.
Buyers should review inspection history, data integrity, staff training, and audit readiness. Modern equipment helps, but incomplete records remain a serious warning.
Product complexity, review questions, and missing evidence can extend the process. The system is not perfectly smooth. Speed cannot replace evidence.
Clinical trials may exclude patients with unusual conditions or multiple illnesses. Ongoing monitoring can detect unexpected problems in everyday use. No study is flawless.
China’s pharmaceutical approval system is designed to protect public health through rigorous oversight by national and specialized regulatory authorities. Before a new drug can enter the market, developers must complete structured preclinical studies and carefully controlled clinical trials. These stages generate evidence about the product’s safety, effectiveness, appropriate dosage, and potential risks. Regulators also examine manufacturing processes, quality controls, stability, and consistency to ensure that every batch meets established standards.
Why is drug approval process strict? Medicines directly affect patients’ health, so approval decisions must be based on reliable scientific evidence rather than commercial expectations. Manufacturing facilities are inspected to verify compliance with Good Manufacturing Practice requirements, including clean operations, qualified personnel, accurate records, and effective quality management. These demanding rules encourage Chinese pharmaceutical suppliers to strengthen research capabilities, improve production systems, maintain transparent documentation, and adopt continuous quality improvement. As a result, suppliers that meet regulatory expectations are better positioned to provide dependable medicines and support long-term confidence in the pharmaceutical sector.
SJ medical